

2.13.3. Descriptor Definitions
Good descriptors make good models. Most of the ones we use are standard in the chemometrics and modeling literature: detailed descriptions can be found in Todeschini and Consonni's comprehensive Handbook of Molecular Descriptors. Most models only make use of molecular descriptors that are functions of the overall molecular structure (e.g., bond and ring counts or connectivity indices) or simple functions of atomic descriptor vectors for their constituent atoms (sums, averages, maxima, minima, correlation vector elements, etc.). S+pKa, the various CYP site of metabolism models and most CYP kinetics models are exceptions in that they do rely on atomic descriptors.
Atomic descriptors include attributes like elemental type, hybridization state, and hydrogen bond donor or acceptor status, but they also include some specific environmental indicator such as topological proximity to acidic or basic groups. Two sets of descriptors developed by Hall and Kier (1991; 1995) have found particularly wide application. One is the intrinsic state I calculated from a base electronegativity and the atom's molecular connectivity. The other is its electrotopological index (E-State) S, which is its intrinsic state perturbed incrementally by the intrinsic states of atoms to which it is connected, down-weighted by the topological distance to each.
Note that the E-state for an an atom type not found in a molecule is set to 0.
In addition to these standard atomic descriptors, we use an advanced set of proprietary atomic charge and reactivity descriptors derived from an analysis of ~1000 diverse multifunctional molecules, 80% of which were used to train our partial atomic charge models and 20% of which served as a test set. GAMESS calculations were run at the B3LYP/6-311G(d,p) level followed by natural population analysis (NPA) to extract atomic atomic charges. A special protocol using natural bond orbitals (NBO) was employed to extract separate sigma and pi components of the partial atomic charges.
The sigma charges were provided as inputs to parameterize a 2D variant (i.e., one based on topological distances) of the electronegativity equalization method (EEM) based on the formalism of Chaves et al (2006) for the EEM kernel. EEM electronegativity and hardness parameters were fit to the ca. 4000 atoms in our training set using 25 hardness and 49 electronegativity parameters using our extension of Chaves's model. To deal with resonance effects in pi systems, a Hückel Model treatment for pi charges was also developed with additional parameters for pi electronegativity, bond resonance and hyperconjugation.
The final model yielded an RMSE 0.048 for sigma charges and 0.040 for pi charges (in atomic units) on the test set. The overall partial atomic charge RMSE is 0.049.
EEM sigma and Hückel pi Fukui indices - derivatives of atomic partial charge with respect to overall electron count - serve as additional atomic descriptors. They represent measures of reactivity in the sigma and pi systems. The pi Fukui indices come in positive and negative "flavors"; f(+) indices relate to pi electron density for the lowest unoccupied molecular orbital (LUMO), whereas f(-) indices relate to pi electron density in the HOMO (Chattaraj 2006; Anderson 2007).
Inclusion of the new EEM-Hückel model charges and Fukui indices improved the performance of most models examined, typically reducing the test set RMSE (or false rate for classification models) by 10-20%.
Molecular descriptors
ADMET Predictor generates 396 molecular descriptors from 2D structures and an additional 50 from 3D structures. Some of these descriptors are used in the mathematical models for the predicted ADMET properties. A few descriptors are provided for free, but access to the full complement is contingent upon having purchased a license for ADMET Modeler or for one of the four ADMET Properties modules. The complete list of descriptors is presented below, along with a brief explanation of each descriptor.
The descriptors are grouped as they are within the Attribute quick selection dialog (1). That dialog is launched by clicking on any of the quick select ions found on the Arrange spreadsheet columns dialog (as well as on many others in ADMET Predictor), which is itself can be launched from the keyboard (Ctrl+Alt+V) or from the menu bar: VIEW >> Arrange Columns...
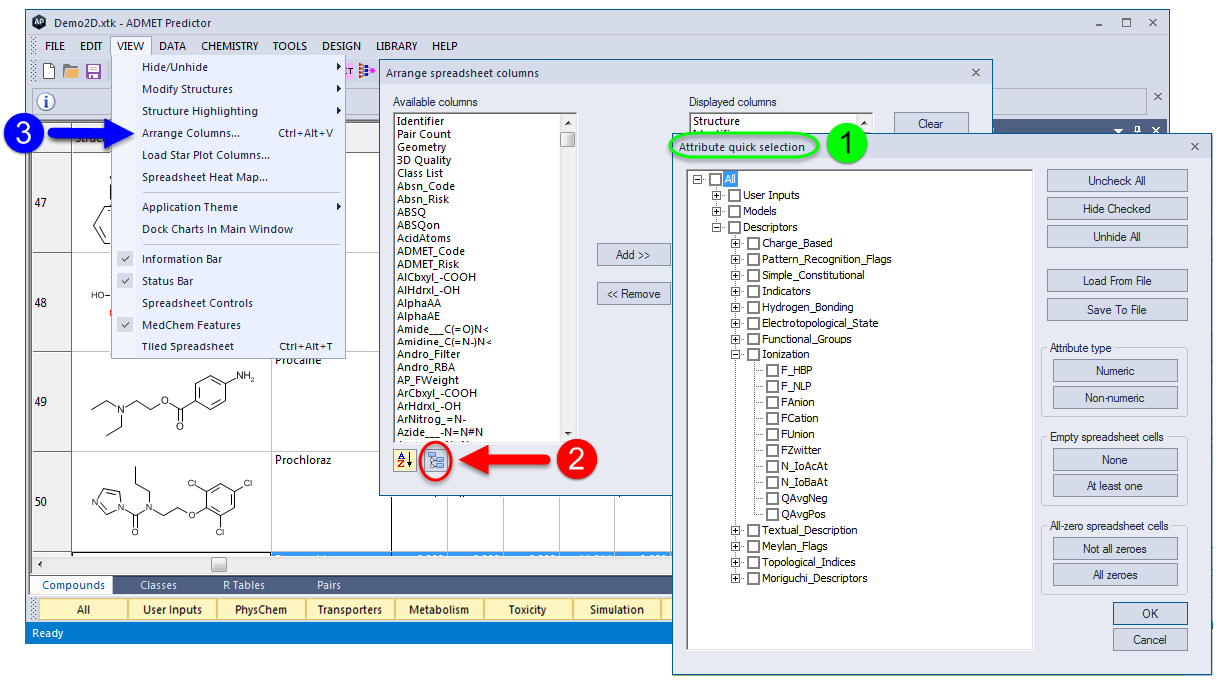
Textual Description descriptors are non-numeric and cannot be used for modeling. Indicators are not intended for modeling; their main purpose is to provide you with notice that the corresponding molecular structure cannot be parsed properly by ADMET Predictor. This may be because it contains metals or other elements not generally encountered in pharmacological chemistry or because it contains problematic atom types.
Constitutional descriptors capture various aspects of molecular size and uniformity of composition, whereas Topological Indices summarize molecular shape. Besides such classical connectivity indices as the Kappa Kier-Hall shape descriptors and the Wiener Index, this group includes topological analogs of 3D shape descriptors. The way T_Radmax is calculated, for example, parallels the way RadMax_3D is calculated. The difference is that the former is applied to the topological distance (bond separation) matrix and the latter is applied to the Euclidean distance matrix. Version 13 of the program introduced descriptors of enantiomeric chirality for the first time (Chirxxx). These will be zero for input structures without chiral indicators (e.g., @ and @@ in SMILES).
Electrotopological State descriptors were devised by Kier and Hall and have been widely used in QSAR applications (see Roy 2012 for a review). Descriptors in the Charge-based group are aggregate functions of the atomic descriptors described above. Ionization descriptors, on the other hand, are derived from our S+pKa microstate analysis. These three descriptors groups are particularly important to many of the models that we develop and distribute.
Textual Description:
|
Formula |
Molecular Formula |
|---|---|
|
AcidAtoms |
Numeric labels and types of all detected acidic atoms |
|
BaseAtoms |
Numeric labels and types of all detected basic atoms |
|
Isoel_pH |
Isoelectric pH, well defined only for zwitterionic/ampholytic compounds. It returns "None" for strict acids, strict bases, neutral compounds. |
Indicators (not generally appropriate for model building):
|
N_Nonorgn |
Number of atoms other than C, N, O, S, P and halogens.* |
|---|---|
|
N_Metal |
Number of metallic element atoms* |
|
N_CYPAtoms |
Number of atoms that can potentially be oxidized by CYPs (excludes epoxidation) |
|
N_CYPSites |
Number of atoms/bonds that can potentially be oxidized by CYPs (includes epoxidation routes) |
|
N_UGTSites |
Number of points of attack by human UGTs |
|
N_AOXSites |
Number of atoms that can potentially be oxidized by Aldehyde Oxidase enzyme |
|
N_Kekule |
Number of Kekule structures per molecule |
|
S_unknown |
The number of unrecognized E-state atom types.* |
|
Unknown_ |
Number of detected but unassigned functional groups.* |
*ADMET Predictor can only properly handle compounds for which N_Nonorgn, N_Metal, S_unknown and Unknown_ are all 0. Compounds for which this is not the case are apt to cause problems in model building and evaluation. Their molecular structures should be checked manually for errors before proceeding. They should probably be set aside, since predictions made for them may not be reliable.
Simple Constitutional Descriptors:
|
MWt |
Molecular Weight (g/mol) |
|---|---|
|
MolVol |
Liquid molal volume (cm3/mol) at the normal boiling point; based on Schroeder's method |
|
VMcGowan |
Mc Gowan's characteristic volume |
|
N_Atoms |
Number of atoms |
|
N_Carbon |
Number of carbons |
|
N_Hydrgn |
Number of hydrogens |
|
N_Ntrgen |
Number of nitrogens |
|
N_Oxygen |
Number of oxygens |
|
N_Phsphr |
Number of phosphorus atoms |
|
N_Sulfur |
Number of sulfurs |
|
N_Boron |
Number of borons |
|
N_Fluorn |
Number of fluorines |
|
N_Chlorn |
Number of chlorines |
|
N_Bromin |
Number of bromines |
|
N_Iodine |
Number of iodines |
|
N_Halogen |
Number of halogen atoms |
|
F_sp3_C |
Fraction of carbon atoms with sp3 hybridization |
|
N_Nonorgn |
Number of "non-organic" atoms (i.e., atoms other than C,N,O,S,P,halogens) |
|
N_Metal |
Number of metallic element atoms |
|
N_Bonds |
Number of bonds |
|
N_FrRotB |
Number of freely rotatable bonds, terminal bonds excluded. At least one bonded atom must be sp3-hybridized. |
|
F_SgleB |
Single bonds as fraction of total bonds. |
|
F_DbleB |
Double bonds as fraction of total bonds. |
|
F_TpleB |
Triple bonds as fraction of total bonds. |
|
F_AromB |
Aromatic bonds as fraction of total bonds. |
|
F_AFRBWF |
Average value of the freely rotatable bond weight factor. The FRBW factor is defined as the minimal atom count fraction per bond. |
|
N_Rings |
Number of rings |
|
N_AromR |
Number of aromatic rings |
|
N_AliphR |
Number of aliphatic rings |
|
N_AntiArom |
Number of Anti-aromatic rings (rings containing all sp2 hybridized atoms with 4N pi electrons and number of pi electrons >= number of ring atoms) |
|
N_Pisyms |
Number of distinct π systems, excluding lone pairs |
|
N_PiSysAtoms |
Total number of atoms involved in pi systems |
|
N_IsolLP |
Number of isolated lone electron pairs |
|
Herndon |
Herndon resonance energy |
Topological Indices:
|
X0 |
Zeroth Order Simple Connectivity Index |
|---|---|
|
X1 |
First Order Simple Connectivity Index |
|
X2 |
Second Order Simple Connectivity Index |
|
Kappa1 |
First Order Simple Kier-Hall shape descriptor |
|
Kappa2 |
Second Order Simple Kier-Hall shape descriptor |
|
Kappa3 |
Third Order Simple Kier-Hall shape descriptor |
|
T_Rgrav |
Topological equivalent of RgGrav_3D |
|
T_Rgeom |
Topological equivalent of RgGeom_3D |
|
T_Radmax |
Topological equivalent of RadMax_3D |
|
T_Rada |
Topological equivalent of Rada__3D |
|
T_Radb |
Topological equivalent of Radb__3D |
|
T_Radc |
Topological equivalent of Radc__3D |
|
T_Rads |
Topological equivalent of Rads__3D |
|
T_HydroR |
Topological equivalent of HydroR__3D |
|
T_MIRxx |
Topological equivalent of MIRxx___3D, but without mass weighting |
|
T_MIRyy |
Topological equivalent of MIRyy___3D, but without mass weighting |
|
T_Dipole |
Topological equivalent of Dipole__3D |
|
T_Lipole |
Topological equivalent of Lipole__3D |
|
T_Grav3 |
Topological equivalent of Grav3__3D |
|
T_RDmtr |
Relative topological diameter: maximal topological distance divided by the number of atoms |
|
T_PSA |
Topological polar surface area, as defined by P. Ertl, et al. |
|
Wiener |
Classical Wiener index of the hydrogen-suppressed molecular graph |
|
Blbn_J |
Balaban distance connectivity index J of the hydrogen-suppressed molecular graph |
|
Blbn_MSD |
Balaban mean square distance index of the hydrogen-suppressed molecular graph |
|
ddRepul |
Number of dipole-dipole repulsions between adjacent or nearby NH groups in ring systems |
|
lpRepul |
Number of lone-pair/lone-pair repulsions between adjacent or nearby N or O atoms in ring systems |
|
Zagreb1 |
Zagreb index of the 1st kind |
|
mZagreb |
Modified Zagreb index of the 1st kind |
|
nSPS |
Normalized spacial score (molecule complexity) |
|
ChirVol |
Descriptor of chirality dependent upon atomic McGowan volumes in individual chiral branches |
|
ChirVolP |
Descriptor of chirality dependent upon atomic McGowan volumes in individual chiral branches, scaled by square of distance from chiral center |
|
ChirVolM |
Descriptor of chirality dependent upon atomic McGowan volumes in individual chiral branches, scaled by inverse square of distance from chiral center |
|
ChirChg |
Descriptor of chirality dependent upon partial atomic charges in individual chiral branches |
|
ChirChgP |
Descriptor of chirality dependent upon partial atomic charges in individual chiral branches, scaled by square of distance from chiral center |
|
ChirChgM |
Descriptor of chirality dependent upon partial atomic charges in individual chiral branches, scaled by inverse square of distance from chiral center |
|
ChirACh |
Descriptor of chirality dependent upon absolute partial atomic charges in individual chiral branches |
|
ChirAChP |
Descriptor of chirality dependent upon absolute partial atomic charges in individual chiral branches, scaled by square of distance from chiral center |
|
ChirAChM |
Descriptor of chirality dependent upon absolute partial atomic charges in individual chiral branches, scaled by inverse square of distance from chiral center |
|
ChirPol |
Descriptor of chirality dependent upon atomic Glen polarizability in individual chiral branches |
|
ChirPolP |
Descriptor of chirality dependent upon atomic Glen polarizability in individual chiral branches, scaled by square of distance from chiral center |
|
ChirPolM |
Descriptor of chirality dependent upon atomic Glen polarizability in individual chiral branches, scaled by inverse square of distance from chiral center |
|
ChirHBA |
Descriptor of chirality dependent upon HB acceptor basicities in individual chiral branches |
|
ChirHBAP |
Descriptor of chirality dependent upon HB acceptor basicities in individual chiral branches, scaled by square of distance from chiral center |
|
ChirHBAM |
Descriptor of chirality dependent upon HB acceptor basicities in individual chiral branches, scaled by inverse square of distance from chiral center |
|
ChirHBD |
Descriptor of chirality dependent upon HB donor acidities in individual chiral branches |
|
ChirHBDP |
Descriptor of chirality dependent upon HB donor acidities in individual chiral branches, scaled by square of distance from chiral center |
|
ChirHBDM |
Descriptor of chirality dependent upon HB donor acidities in individual chiral branches, scaled by inverse square of distance from chiral center |
|
ChirFlx |
Descriptor of chirality dependent upon bond flexibilities in individual chiral branches |
|
ChirFlxP |
Descriptor of chirality dependent upon bond flexibilities in individual chiral branches, scaled by square of distance from chiral center |
|
ChirFlxM |
Descriptor of chirality dependent upon bond flexibilities in individual chiral branches, scaled by inverse square of distance from chiral center |
|
BCUT_MW_HI |
Highest eigenvalue of Burden matrix weighted by atomic weights |
|
BCUT_MW_LO |
Lowest eigenvalue of Burden matrix weighted by atomic weights |
|
BCUT_LP_HI |
Highest eigenvalue of Burden matrix weighted by atomic lipophilicities |
|
BCUT_LP_LO |
Lowest eigenvalue of Burden matrix weighted by atomic lipophilicities |
|
lnHosoyaZ |
Logarithm of the sum of the absolute values of the coefficients of the characteristic polynomial of the molecular graph divided by the number of atoms |
|
InfCntnt |
Average information content of the molecular graph |
Atom-type Electropological State Indices
|
SsCH3 |
Atom-type E-state index for -CH3 groups |
|---|---|
|
SdCH2 |
Atom-type E-state index for =CH2 groups |
|
SssCH2 |
Atom-type E-state index for -CH2- groups |
|
StCH |
Atom-type E-state index for #CH groups |
|
SdsCH |
Atom-type E-state index for =CH- groups |
|
SaaCH |
Atom-type E-state index for aCHa groups |
|
SsssCH |
Atom-type E-state index for >CH- groups |
|
SddC |
Atom-type E-state index for =C= groups |
|
StsC |
Atom-type E-state index for #C- groups |
|
SdssC |
Atom-type E-state index for =C< groups |
|
SaasC |
Atom-type E-state index for -Caa groups |
|
SaadC |
Atom-type E-state index for =Caa groups (e.g., C=O in theophylline) |
|
SaaaC |
Atom-type E-state index for aCaa groups |
|
SssssC |
Atom-type E-state index for >C< groups |
|
SsNH2 |
Atom-type E-state index for -NH2 groups |
|
SdNH |
Atom-type E-state index for =NH groups |
|
SssNH |
Atom-type E-state index for -NH- groups |
|
SaaNH |
Atom-type E-state index for aNHa groups |
|
StN |
Atom-type E-state index for #N groups |
|
SdsN |
Atom-type E-state index for =N- groups |
|
SdssN |
Atom-type E-state index for =N< groups (secondary N-oxide) |
|
SdsssN |
Atom-type E-state index for =N<- groups (tertiary N-oxide) |
|
StsN |
Atom-type E-state index for -N+# groups (diazonium) |
|
StdN |
Atom-type E-state index for #N= groups (azide) |
|
SaaN |
Atom-type E-state index for aNa groups |
|
SsssN |
Atom-type E-state index for >N- groups |
|
SddsN |
Atom-type E-state index for -N== groups (nitro) |
|
SaasN |
Atom-type E-state index for aaN- groups (e.g., substituted imidazole) |
|
SaadN |
Atom-type E-state index for aaN= groups (aromatic N-oxide) |
|
SaaaN |
Atom-type E-state index for aaNa groups (N-fused rings) |
|
SssssN |
Atom-type E-state index for >N+< groups (onium) |
|
SsOH |
Atom-type E-state index for -OH groups |
|
SdO |
Atom-type E-state index for =O groups |
|
SssO |
Atom-type E-state index for -O- groups |
|
SsO- |
Atom-type E-state index for coordinated O- groups |
|
SaaO |
Atom-type E-state index for aOa groups |
|
SsPH2 |
Atom-type E-state index for -PH2 groups |
|
SssPH |
Atom-type E-state index for -PH- groups |
|
SdssPH |
Atom-type E-state index for =PH< groups (phosphinate) |
|
SsssP |
Atom-type E-state index for >P- groups |
|
SdsssP |
Atom-type E-state index for ->P= groups |
|
SssssP |
Atom-type E-state index for >P+< groups (phosphonium) |
|
SsssssP |
Atom-type E-state index for ->P< groups |
|
SsSH |
Atom-type E-state index for -SH groups |
|
SdS |
Atom-type E-state index for =S groups |
|
SssS |
Atom-type E-state index for -S- groups |
|
SsssS |
Atom-type E-state index for -S+< groups (sulfonium) |
|
SaaS |
Atom-type E-state index for aSa groups |
|
SdssS |
Atom-type E-state index for >S= groups (sulfone) |
|
SddssS |
Atom-type E-state index for >S== groups (sulfate) |
|
SssssssS |
Atom-type E-state index for ->S<- groups |
|
SsssB |
Atom-type E-state index for -B< groups |
|
SaasB |
Atom-type E-state index for aaB- groups |
|
SsF |
Atom-type E-state index for -F groups |
|
SsCl |
Atom-type E-state index for -Cl groups |
|
SsBr |
Atom-type E-state index for -Br groups |
|
SsI |
Atom-type E-state index for -I groups |
|
SHsOH |
Atom-type hydrogen E-state index for -OH groups |
|
SHdNH |
Atom-type hydrogen E-state index for =NH groups |
|
SHsSH |
Atom-type hydrogen E-state index for -SH groups |
|
SHsNH2 |
Atom-type hydrogen E-state index for -NH2 groups |
|
SHssNH |
Atom-type hydrogen E-state index for -NH- groups |
|
SHtCH |
Atom-type hydrogen E-state index for #CH groups |
|
SHCH_321 |
Atom-type hydrogen E-state index for -CH3, -CH2- and >CH- groups (saturated aliphatic carbon) |
|
SHdCH2 |
Atom-type hydrogen E-state index for =CH2 groups |
|
SHdsCH |
Atom-type hydrogen E-state index for =CH- groups |
|
SHaaCH |
Atom-type hydrogen E-state index for aCHa groups (aromatic carbons) |
Charge-based Descriptors*:
|
FormalQ |
Formal electric charge |
|---|---|
|
ABSQ |
Sum of Absolute Values of PEOE Partial Atomic Charges |
|
ABSQon |
As above, but only on O and N atoms |
|
MaxQ |
Maximal PEOE Partial Atomic Charge |
|
MinQ |
Minimal PEOE Partial Atomic Charge |
|
NPA_ABSQ |
Sum of absolute values of estimated NPA partial atomic charges |
|
NPA_AQon |
As above, but only on O and N atoms |
|
NPA_AQh |
As above, but only on H atoms |
|
NPA_AQc |
As above, but only on C atoms |
|
NPA_MaxQ |
Maximal Estimated NPA Partial Atomic Charge |
|
NPA_MinQ |
Minimal Estimated NPA Partial Atomic Charge |
|
NPA_Q1 |
First component of the autocorrelation vector of estimated NPA partial atomic charges |
|
NPA_Q2 |
Second component of the autocorrelation vector of estimated NPA partial atomic charges |
|
NPA_Q3 |
Third component of the autocorrelation vector of estimated NPA partial atomic charges |
|
NPA_Q4 |
Fourth component of the autocorrelation vector of estimated NPA partial atomic charges |
|
NPA_Q5 |
Fifth component of the autocorrelation vector of estimated NPA partial atomic charges |
|
NPA_Q6 |
Sixth component of the autocorrelation vector of estimated NPA partial atomic charges |
|
EEM_AFon |
Sum of absolute values of sigma Fukui indices on N and O |
|
EEM_AFc |
Sum of absolute values of sigma Fukui indices on C |
|
EEM_AFpl |
Sum of absolute values of sigma Fukui indices on polar atoms |
|
EEM_AFnp |
Sum of absolute values of sigma Fukui indices on nonpolar atoms |
|
EEM_MaxF |
Maximum sigma Fukui index |
|
EEM_XFon |
Maximum sigma Fukui index on N and O |
|
EEM_NFon |
Minimum sigma Fukui index on N and O |
|
EEM_XFh |
Maximum sigma Fukui index on H |
|
EEM_XFc |
Maximum sigma Fukui index on C |
|
EEM_NFc |
Minimum sigma Fukui index on C |
|
EEM_XFpl |
Maximum sigma Fukui index on polar atoms |
|
EEM_NFpl |
Minimum sigma Fukui index on polar atoms |
|
EEM_XFnp |
Maximum sigma Fukui index on nonpolar atoms |
|
EEM_NFnp |
Minimum sigma Fukui index on nonpolar atoms |
|
EEM_F1 |
First component of the autocorrelation vector of sigma Fukui indices |
|
EEM_F2 |
Second component of the autocorrelation vector of sigma Fukui indices |
|
EEM_F3 |
Third component of the autocorrelation vector of sigma Fukui indices |
|
EEM_F4 |
Fourth component of the autocorrelation vector of sigma Fukui indices |
|
EEM_F5 |
Fifth component of the autocorrelation vector of sigma Fukui indices |
|
EEM_F6 |
Sixth component of the autocorrelation vector of sigma Fukui indices |
|
Pi_ABSQ |
Sum of absolute values of Hückel pi atomic charges |
|
Pi_AQo |
As above, but only on O atoms |
|
Pi_AQn |
As above, but only on N atoms |
|
Pi_AQc |
As above, but only on C atoms |
|
Pi_MaxQ |
Maximum Hückel pi atomic charge |
|
Pi_MinQ |
Minimum Hückel pi atomic charge |
|
Pi_Q1 |
First component of the autocorrelation vector of Hückel pi atomic charges |
|
Pi_Q2 |
Second component of the autocorrelation vector of Hückel pi atomic charges |
|
Pi_Q3 |
Third component of the autocorrelation vector of Hückel pi atomic charges |
|
Pi_Q4 |
Fourth component of the autocorrelation vector of Hückel pi atomic charges |
|
Pi_Q5 |
Fifth component of the autocorrelation vector of Hückel pi atomic charges |
|
Pi_Q6 |
Sixth component of the autocorrelation vector of Hückel pi atomic charges |
|
Pi_AFPlc |
Sum of absolute values of pi Fukui(+) indices on carbon |
|
Pi_AFMic |
Sum of absolute values of pi Fukui(-) indices on carbon |
|
Pi_MaxFPl |
Maximum pi Fukui(+) index |
|
Pi_MaxFMi |
Maximum pi Fukui(-) index |
|
Pi_FPl1 |
First component of the autocorrelation vector of pi Fukui(+) indices |
|
Pi_FPl2 |
Second component of the autocorrelation vector of pi Fukui(+) indices |
|
Pi_FPl3 |
Third component of the autocorrelation vector of pi Fukui(+) indices |
|
Pi_FPl4 |
Fourth component of the autocorrelation vector of pi Fukui(+) indices |
|
Pi_FPl5 |
Fifth component of the autocorrelation vector of pi Fukui(+) indices |
|
Pi_FPl6 |
Sixth component of the autocorrelation vector of pi Fukui(+) indices |
|
Pi_FMi1 |
First component of the autocorrelation vector of pi Fukui(-) indices |
|
Pi_FMi2 |
Second component of the autocorrelation vector of pi Fukui(-) indices |
|
Pi_FMi3 |
Third component of the autocorrelation vector of pi Fukui(-) indices |
|
Pi_FMi4 |
Fourth component of the autocorrelation vector of pi Fukui(-) indices |
|
Pi_FMi5 |
Fifth component of the autocorrelation vector of pi Fukui(-) indices |
|
Pi_FMi6 |
Sixth component of the autocorrelation vector of pi Fukui(-) indices |
|
EqualChi |
Equalized molecular electronegativity |
|
EqualEta |
Equalized molecular hardness |
|
Elephity |
Electrophilicity index |
|
N_Electr |
Total number of electrons in a molecule |
|
PolarizG |
Polarizability in A3 calculated by Glen's method |
|
PolarizM |
Polarizability in A3 calculated by Miller's method |
** *** The charge unit is the absolute charge on an electron.
Hydrogen Bonding Descriptors:
|
IHB |
Number of Intra-molecular Hydrogen Bonds |
|---|---|
|
PHB |
Proximity Effects of HB Donors and HB Acceptors. Similar to IHB, but counts all possible donor-acceptor pairs within the IHB range. |
|
HBD |
Number of OH and NH Hydrogen Bond Donors |
|
HBDH |
Number of OH and NH Hydrogen Bond Donor Protons |
|
HBA |
Number of O and N Hydrogen Bond Acceptors |
|
HBDo |
Number of Oxygen-based Hydrogen Bond Donors |
|
HBAo |
Number of Oxygen-based Hydrogen Bond Acceptors |
|
HBDn |
Number of Nitrogen-based Hydrogen Bond Donors |
|
HBAn |
Number of Nitrogen-based Hydrogen Bond Acceptors |
|
HBDch |
Sum of Estimated NPA Partial Atomic Charges on HB Donor Hydrogens |
|
HBAch |
Sum of Estimated NPA Partial Atomic Charges on HB Acceptors |
|
HBDoch |
Sum of Estimated NPA Partial Atomic Charges on Oxygen-based HB Donor Hydrogens |
|
HBAoch |
Sum of Estimated NPA Partial Atomic Charges on Oxygen-based HB Acceptors |
|
HBDnch |
Sum of Estimated NPA Partial Atomic Charges on Nitrogen-based HB Donor Hydrogens |
|
HBAnch |
Sum of Estimated NPA Partial Atomic Charges on Nitrogen-based HB Acceptors |
|
lrIHB |
Number of long-range (>3 bonds away) intramolecular hydrogen donor-acceptor pairs |
|
avgIHBdist |
Average topological distance between long-range intramolecular hydrogen donor-acceptor pairs |
|
avgIHBstrn |
Average strength of interaction (HBacidity*HBbasicity) between long-range intramolecular hydrogen donor-acceptor pairs |
|
maxIHBstrn |
Maximum strength of interaction (HBacidity*HBbasicity/100) between long-range intramolecular hydrogen donor-acceptor pairs |
|
distIHBmax |
Topological distance between strongest long-range intramolecular hydrogen donor-acceptor pair |
|
avgHBAperD |
Average number of long-range intramolecular hydrogen acceptors per donor |
|
avgHBDperA |
Average number of long-range intramolecular hydrogen donors per acceptor |
Molecular Ionization Descriptors:
|
N_IoAcAt |
Number of recognized ionizable atom types that are acidic |
|---|---|
|
N_IoBaAt |
Number of recognized ionizable atom types that are basic |
|
FAnion |
Cumulative Contribution of Purely Anionic Species to Fraction Ionized at Specified pH (default = 7.4) |
|
FCation |
Cumulative Contribution of Purely Cationic Species to Fraction Ionized at Specified pH (default = 7.4) |
|
FUnion |
Cumulative Contribution of All Species with Zero Formal Charge to Fraction Ionized at Specified pH (default = 7.4) |
|
FZwitter |
Portion of FUnion Contributed by Zwitterionic Species independent of pH |
|
QAvgNeg |
Absolute Value of the Population Average Across All Ionized Species of the Net Formal Negative Charge Calculated at Specified pH (default = 7.4) |
|
QAvgPos |
Population Average Across All Ionized Species of the Net Formal Positive Charge Calculated at Specified pH (default = 7.4) |
|
F_NLP |
Population Average Across All Ionized Species of the Number of Lone Electron Pairs on N,O,S,P, Divided by the Number of Atoms |
|
F_HBP |
Population Average Across All Ionized Species of the Number of Protons Available for Hydrogen Bonding, Divided by the Number of Atoms |
Functional Groups:
|
AlHdrxl_-OH |
Number of aliphatic hydroxyl groups |
|---|---|
|
ArHdrxl_-OH |
Number of aromatic hydroxyl groups |
|
Carbonyl_C=O |
Number of carbonyl (ketone or aldehyde) groups |
|
Ether___-O- |
Number of ether groups |
|
PriAmine_-NH2 |
Number of primary amines |
|
PriAmAli_-NH2 |
Number of primary amines attached to aliphatic carbon |
|
PriAmAro_-NH2 |
Number of primary amines attached to aromatic carbon |
|
SecAmine_>NH |
Number of primary and aliphatic N secondary amines |
|
TerAmine_>N- |
Number of tertiary amine groups |
|
QuaAmine_>[N+]< |
Number of quaternary amine groups |
|
ArNitrog_=N- |
Number of aromatic nitrogens |
|
Imine___=N- |
Number of Imine groups |
|
Nitrile_C#N |
Number of Nitrile groups |
|
Isocynd_[N+]#[C-] |
Number of Isocyanide groups |
|
Thiol___-SH |
Number of Thiol groups |
|
ThioCrbl_C=S |
Number of Thiocarbonyl groups |
|
Sulfide_-S- |
Number of Sulfide groups |
|
Slfonium_>[S+]- |
Number of Sulfonium groups |
|
Phsphine_>P- |
Number of Phosphine groups |
|
HydxlAmn_>N-O- |
Number of Hydroxyamine groups |
|
Oxime___=N-O- |
Number of Oxime groups |
|
Nitroso_-N=O |
Number of Nitroso groups |
|
N-oxide_>[N+][O-] |
Number of N-oxide groups |
|
Hdrzine_>N-N< |
Number of Hydrazine groups |
|
Hdrzone_=N-N< |
Number of Hydrazone groups |
|
Azo_____-N=N- |
Number of Azo groups |
|
Diazo___-[N+]#N |
Number of Diazo groups |
|
ThioAmin_>N-S- |
Number of Thioamine groups |
|
Disulfde_-S-S- |
Number of Disulfide groups |
|
Sulfoxde_>S=O |
Number of Sulfoxide groups |
|
Thioxime_=N-S- |
Number of Thioxime groups |
|
Nitro___-NO2 |
Number of Nitro groups |
|
Nitrite_-O-N=O |
Number of Nitrite groups |
|
Ntrosam_>N-N=O |
Number of Nitrosamine groups |
|
Azoxy___-N(=O)=N- |
Number of Azoxy groups |
|
Azide___-N=N#N |
Number of Azide groups |
|
Sulfone_>S(=O)=O |
Number of Sulfone groups |
|
Sulfinat_S(=O)O- |
Number of Sulfinate groups |
|
Phsporat_P(=O)O- |
Number of Phosphorate groups |
|
Triazo__-N-N-N- |
Number of Triazo groups |
|
Oxadiazo_-N-O-N- |
Number of Oxadiazo groups |
|
Thiadiaz_-N-S-N- |
Number of Thiadiazo groups |
|
Triazene_-N=N-N- |
Number of Triazene groups |
|
Nitrate_-O-NO2 |
Number of Nitrate groups |
|
NNitro__>N-NO2 |
Number of N-connected nitro groups |
|
Sulfonat_-SO3- |
Number of Sulfonate groups |
|
Sulfite_-SO3< |
Number of Sulfite groups |
|
Sulfonmd_-SO2-N< |
Number of Sulfonamide groups |
|
PriSlfmd_-SO2-NH2 |
Number of Primary Sulfonamide groups |
|
Phsponat_-PO3< |
Number of Phosphonate groups |
|
Sphspnat_-SPO2< |
Number of Thiolophosphonate groups |
|
Phspite_-OPO2< |
Number of Phosphite groups |
|
Tetrazo_-NNNN- |
Number of Tetrazo groups |
|
Oxdzoxde_-N(=O)ON- |
Number of Oxadiazooxide groups |
|
NOHydxam_
|
Number of N-nitrosohydroxylamine groups |
|
Sulfamid_>NSO2N< |
Number of Sulfamide groups |
|
Sulfate_-OSO3 |
Number of Sulfate groups |
|
ThioSulf_-SSO3 |
Number of Thiosulfate groups |
|
Phspate_O=PO3<- |
Number of Phosphate groups |
|
PhspatS_
|
Number of ThioloPhosphate groups: O=P(X)(X)X where X = S,O |
|
Sphspat_S=PO3<- |
Number of ThioPhosphate groups |
|
SphspatS_
|
Number of ThioThioloPhosphate groups: S=P(X)(X)X where X = S,O |
|
PhspatN_
|
Number of Phosphamide groups: O=P(X)(X)X where X = N,O |
|
SphspatN_
|
Number of ThioPhosphamide groups: S=P(X)(X)X where X = N,O |
|
PhsptSN_
|
Number of ThioloPhosphamide groups: O=P(X)(X)X where (X = N,O,S) |
|
SPhsptSN_
|
Number of ThioThioloPhosphamide groups: S=P(X)(X)X where (X = N,O,S) |
|
Diphspat_
|
Number of Diphosphate groups |
|
Triphspt_
|
Number of Triphosphate groups |
|
AlCbxyl_-COOH |
Number of Aliphatic Carboxyl groups |
|
ArCbxyl_-COOH |
Number of Aromatic Carboxyl groups |
|
Ester___C(=O)OC |
Number of Ester groups |
|
Amide___C(=O)N< |
Number of Amide groups |
|
Thioamd_C(=S)N< |
Number of Thioamide groups |
|
Amidine_C(=N-)N< |
Number of Amidine groups |
|
Isocyant_-N=C=O |
Number of Isocyanate groups |
|
Thiocynt_-S-C#N |
Number of Thiocyanate groups |
|
Isothcnt_-N=C=S |
Number of Isothiocyanate groups |
|
Urea____>NC(=O)N< |
Number of Urea groups |
|
Carbmte_>NC(=O)O- |
Number of Carbamate groups |
|
Guandne_>NC(=N-)N< |
Number of Guanidine groups |
|
Imide___O=C(N-)C=O |
Number of Imide groups |
|
Barbitur_
|
Number of Barbiturate groups |
|
AromAmide_c(=O)[nH] |
Number of aromatic amide bonds (e.g. as in 2-pyridone) |
Moriguchi Descriptors used in calculating MlogP:
|
Parameter |
Description |
Example |
|---|---|---|
|
M_CX |
Summation of numbers of carbon and halogen atoms weighted by C:1.0, F:0.5, Cl:1.0, Br:1.5, and I:2.0. |
|
|
M_NO |
Total number of N and O atoms |
|
|
M_PRX |
Proximity effect of N & O; where X-Y:2.0, X-A-Y:1.0 (X,Y can be N or O and A can be C,S, or P) with a correction (-1) for carboxamide/sulfonamide. |
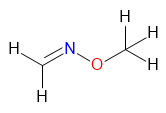
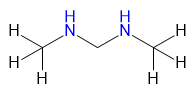
= 1 |
|
M_UB |
Total number of unsaturated bonds except those in NO2 |

= 3 |
|
M_HB |
Number of intramolecular hydrogen bonds as ortho –OH and –C(=O)-R, or –OH and –NH2, or –NH2 and –COOH, or 8-OH/NH2 in quinolines, etc. 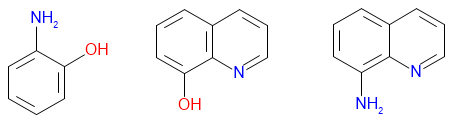
|
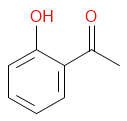
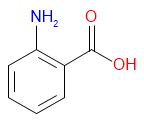
|
|
M_POL |
Number of aromatic polar substituents (aromatic substituents excluding Ar-CX2- and Ar-CX=C<, where X: is C or H. |
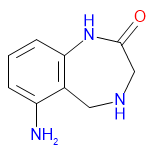
|
|
M_AMP |
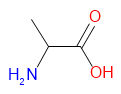
Amphoteric property; α-amino acid:1.0, amino benzoic acid: 0.5, pyridinecarboxylic acid 0.5. 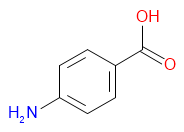
=0.5 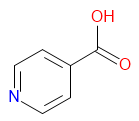
=0.5 |
|
|
M_ALK |
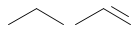
Indicator variable for alkane, alkene, cycloalkane, or cycloalkene (hydrocarbons with 0 or 1 double bond) 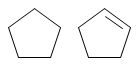
|
|
|
M_RNG |
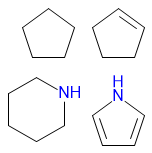
Indicator variable for the presence of ring structures except benzene and its condensed rings (aromatic, heteroaromatic, and hydrocarbon rings)- e.g., excluding: 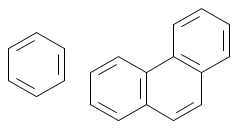
|
|
|
M_QN |
Quaternary nitrogen: >N+<: 1.0, N oxide:0.5 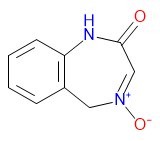
=0.5 |

|
|
M_NO2 |
Number of nitro groups |
|
|
M_NCS |
Isothiocyanate (-N=C=S): 1.0, thiocyanate (-S-C#N): 0.5 |
|
|
M_BLM |
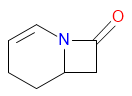
Indicator variable for the presence of β-lactam. |
|
Pattern Recognition Flags:
|
AlphaAA |
Alpha Amino Acid Flag |
|
PrAlphAA |
Primary Alpha Amino Acid Flag |
|
AlphaAE |
Alpha Amino Ester Flag |
|
PrAlphAE |
Primary Alpha Amino Ester Flag |
|
Steroid |
Flag Indicating Steroid-like Fused Ring Subunit |
|
N_Thiophenes |
Number of thiophene rings |
|
N_Thiazoles |
Number of thiazole rings |
|
N_Furans |
Number of furan rings |
|
N_Oxazoles |
Number of oxazole rings |
|
N_Imidazoles |
Number of imidazole rings |
|
N_Piperidines |
Number of piperidine rings |
|
N_Piperazines |
Number of piperazine rings |
|
N_Pyridines |
Number of pyridine rings |
|
N_PhenolOH |
Number of -OH groups connected to phenyl rings |
|
N_Ureas |
Number of urea moieties (note: this is different from Urea functional groups!) |
|
N_UnbrChains |
Number of unbranched chains of sp3 atoms of length >= 4 |
|
N_QuaternaryN |
Number of aliphatic quaternary amines (QuaAmine_>[N+]< also counts aromatic q.a.) |
|
N_Sulfide |
Number of aliphatic sulfides (Sulfide_-S- also counts aromatic sulfides) |
|
N_PrimAmides |
Number of primary amides (note: this is different from Amide functional groups!) |
|
Macrocycle |
An indicator of the presence of a large (12 atoms or more) ring |
Indicator variables (not counts) specific to Meylan solubility models:
|
Parameter |
Description |
Exceptions |
|---|---|---|
|
H_AlAlco
|
Compounds with exactly one OH group attached to an aliphatic hydrocarbon

|
Compounds with any functional group, including a second OH group 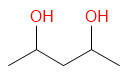
|
|
H_AlAcid
|
Compounds with one or more carboxylate groups attached to aliphatic carbons 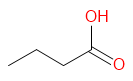
|
Amino acids (aliphatic or aromatic amines, except quaternary) and amides 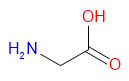
|
|
H_AlAmin
|
Compounds with primary, secondary, or tertiary amines attached to aliphatic carbons 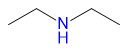

|
Compounds with a melting point greater > 25ºC |
|
H_ArAcid
|
Compounds with at least one carboxylate group attached to an aromatic carbon 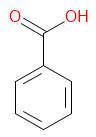
|
Compounds with an amino group attached to the same ring to which the carboxylate is attached. “Amino” = aliphatic nitrogen whose only attachments are carbons or other nitrogens. 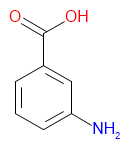
|
|
H_Phenol
|
Compounds with at least one OH attached to an aromatic carbon 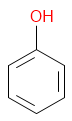
|
Compounds with an amino group on the same ring and those with nitro, alkyloxy or other OH ortho to OH 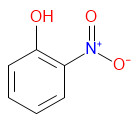
|
|
H_AlPyri
|
Compounds with at least one pyridine ring 

|
Only alkyl substituents allowed; carbon attached to ring must by CH3,
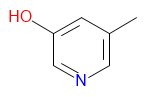
|
|
H_Azo
|
Compounds with -N=N- |
Both N’s must be attached to carbon |
|
H_Nitrile
|
Compounds with -CN |
N=C-CN, C=N-CN |
|
H_Hcarb
|
Aliphatic compounds with only C and H 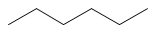
|
|
|
H_Nitro
|
Aliphatic and aromatic compounds with nitro (-NO2) group 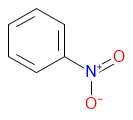
|
N attachment or any amino or OH group if the nitro group is on an aromatic ring 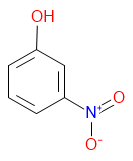
|
|
H_SO2
|
Compounds with at least one sulfonamide on aromatic ring plus an other substituent that is ketone, sulfone, or another sulfonamide on the same ring; also applies to compounds with -S(=O)-C-C(=O)-C (number of Os on S is not limited) 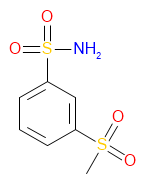
|
|
|
H_Falkan
|
Hydrocarbons with at least one fluorine 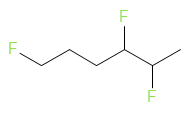
|
|
|
H_PAH
|
Hydrocarbons with at least three rings, at least two of which must be aromatic. 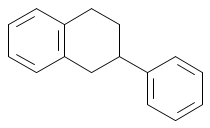
|
|
|
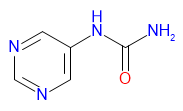
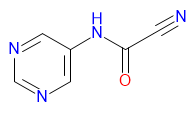
H_MultiN
|
Compounds with at least one of the following:
|
Compounds with at least one of the following:
|
|
H_AmAcid
|
Compound contains at least one carboxylate and at least one amino group. |
Compounds with aromatic attachment of at least one carboxylate group. |
3D Descriptors (length unit=angstrom, mass unit=dalton):
|
RgGrav__3D |
Gravitational radius of gyration |
|---|---|
|
RgGeom__3D |
Geometrical radius of gyration |
|
RadMax__3D |
Distance between center of mass and the most distant atom |
|
Propr1__3D |
Simulations Plus proprietary descriptor |
|
Rada__3D |
Largest ellipsoidal radius of the molecule, based on geometry |
|
Radb__3D |
Medium ellipsoidal radius of the molecule, based on geometry |
|
Radc__3D |
Smallest ellipsoidal radius of the molecule, based on geometry |
|
Rads__3D |
Projected geometrical radius of the molecule, averaged over all orientations |
|
HydroR__3D |
Hydrodynamic radius of molecular ellipsoid |
|
MIRxx___3D |
Largest relative principal moment of inertia |
|
MIRyy___3D |
Medium relative principal moment of inertia |
|
MIRzz___3D |
Smallest relative principal moment of inertia |
|
SM2xx___3D |
Smallest relative principal static moment of the second order |
|
SM2yy___3D |
Medium relative principal static moment of the second order |
|
SM2zz___3D |
Largest relative principal static moment of the second order |
|
BoxX____3D |
Smallest dimension of the tightest enclosing box in A |
|
BoxY____3D |
Medium dimension of the tightest enclosing box in A |
|
BoxZ____3D |
Largest dimension of the tightest enclosing box in A |
|
DStokes_3D |
Stokes-Einstein diffusion coefficient (cm^2/s x 10^5) |
|
Propr2__3D |
Simulations Plus proprietary descriptor |
|
Grav3___3D |
Cube root of Wessel gravitational index |
|
TotASA__3D |
Total solvent accessible surface area in A^2 |
|
PolASA__3D |
Polar solvent accessible surface area in A^2 |
|
NpoASA__3D |
Nonpolar solvent accessible surface area in A^2 |
|
SolvE___3D |
Aqueous solvation energy in kcal/mol |
|
SolvEMt_3D |
Length of the solvation energy moment vector in A*kcal/mol |
|
PEoED___3D |
Proximity effects of electron donors. Descriptor derived from Anna Seelig's work on P-glycoprotein substrate recognition patterns. |
|
PEoEDIa_3D |
Proximity effects of electron donors of type I including atoms with hydrogens. Descriptor derived from Anna Seelig's work on P-glycoprotein substrate recognition patterns. |
|
PEoEDIb_3D |
Proximity effects of electron donors of type I excluding atoms with hydrogens. Descriptor derived from Anna Seelig's work on P-glycoprotein substrate recognition patterns. |
|
PEoEDIIa3D |
Proximity effects of electron donors of type II including atoms with hydrogens. Descriptor derived from Anna Seelig's work on P-glycoprotein substrate recognition patterns. |
|
PEoEDIIb3D |
Proximity effects of electron donors of type II excluding atoms with hydrogens. Descriptor derived from Anna Seelig's work on P-glycoprotein substrate recognition patterns. |
|
HBAsa___3D |
Total solvent accessible surface area on HB acceptors |
|
HBAwsa__3D |
Estimated NPA charge-weighted solvent accessible surface area on HB acceptors |
|
PosASA__3D |
Positively NPA-charged solvent accessible surface area |
|
NegASA__3D |
Negatively NPA-charged solvent accessible surface area |
|
Dipole__3D |
Estimated NPA dipole moment |
|
Lipole__3D |
3D lipole moment (an estimate of the separation of hydrophobic and hydrophilic regions of a molecule much like dipole moment measures the separation of positive and negative charge). |
|
WHIM_L1s_3D |
1st component size directional WHIM index / weighted by relative I-states |
|
WHIM_L2s_3D |
2nd component size directional WHIM index / weighted by relative I-states |
|
WHIM_L3s_3D |
3rd component size directional WHIM index / weighted by relative I-states |
|
WHIM_E1s_3D |
1st component accessibility directional WHIM index / weighted by relative I-states |
|
WHIM_E2s_3D |
2nd component accessibility directional WHIM index / weighted by relative I-states |
|
WHIM_E3s_3D |
3rd component accessibility directional WHIM index / weighted by relative I-states |
|
WHIM_As_3D |
2nd order non-directional WHIM index / weighted by relative I-states |
|
WHIM_Vs_3D |
3rd order non-directional WHIM index / weighted by relative I-states |
|
WHIM_Ks_3D |
Non-directional WHIM shape index / weighted by relative I-states |
|
Eccentricity_3D |
Eccentricity of inertia |
|
InertialSF_3D |
Inertial shape factor |
|
MIratio_yx_3D |
Ratio of principal moments of inertia yy/xx |
|
MIratio_zx_3D |
Ratio of principal moments of inertia zz/xx |